

Intake
- Call center, email, web form and field service
- App store reviews and support tickets for SaMD
- Required complaint fields enforced
- Aware date captured on receipt


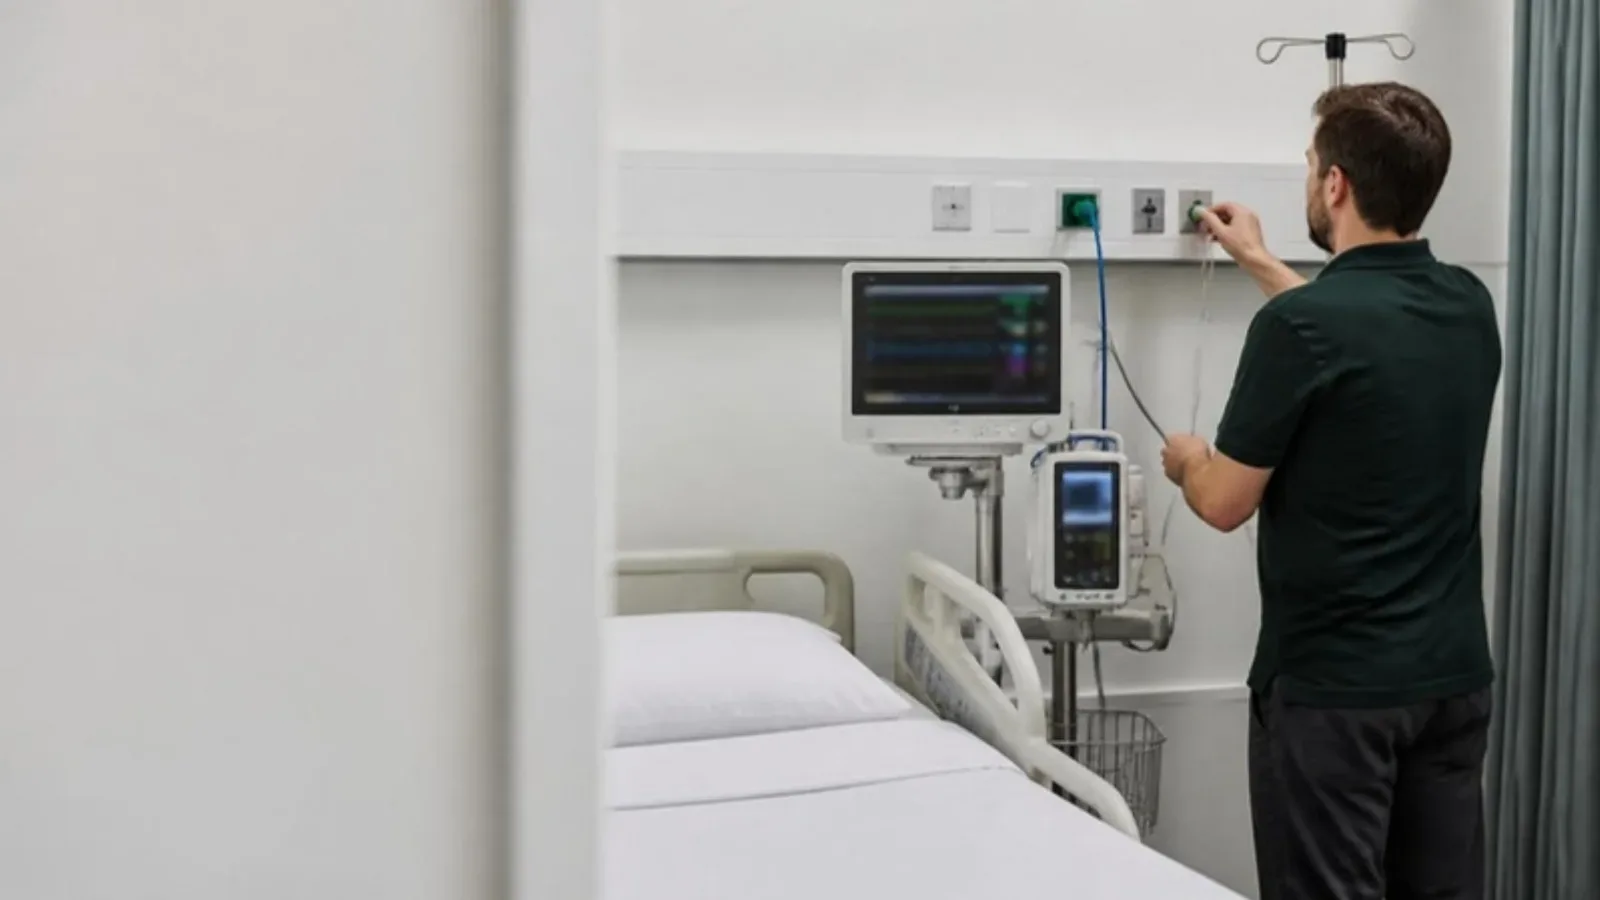
A complaint that sits in an inbox can turn into a late medical device report. Agnotic Technologies builds complaint handling software that takes every complaint from intake to a documented reportability decision and an eMDR filing, with AI that helps sort complaints while your people make every call.
Trusted by global innovators











Complaint handling
Medical device complaint handling software takes each complaint from intake through investigation, the MDR reportability decision and electronic submission to FDA, and keeps the records FDA expects. Agnotic Technologies builds complaint handling software for device and SaMD makers, using AI only to sort and group complaints while qualified people make every reportability call.
The clock starts when anyone in your company becomes aware of a possible reportable event, not when the complaint reaches the quality team. So the software is built around the aware date.
Complaints also feed corrective action and trend analysis. Those live in your quality management system; this software hands each complaint over with the link intact.
Decision table
The table the reportability screen is built from, per 21 CFR Part 803 and the QMSR. Your regulatory lead confirms the wording and owns every decision.
| Complaint event | Report it triggers | Clock | Record the software keeps |
|---|---|---|---|
| Device may have caused or contributed to a death | Medical device report to FDA | 30 calendar days from becoming aware | Event details, investigation, reportability reasoning, submission and FDA acknowledgment |
| Device may have caused or contributed to a serious injury | Medical device report to FDA | 30 calendar days from becoming aware | The same, plus which serious injury criteria were met |
| Malfunction that would likely cause death or serious injury if it recurred | Medical device report to FDA | 30 calendar days from becoming aware | Malfunction analysis and the recurrence assessment |
| Reportable event that needs remedial action to prevent an unreasonable risk of substantial harm | 5-day report | 5 work days from becoming aware | The remedial action decision and any linked correction or removal record |
| Event type covered by a written FDA request | 5-day reports for those events | 5 work days, for the period FDA names | The request, its period and every report filed under it |
| New information after a report was filed | Supplemental report | 30 calendar days from receiving it | Report version history and what changed |
| Complaint with no possible adverse event | No medical device report | None | Complaint record, investigation or the reason none was needed, and the not-reportable decision |
A correction or removal started to reduce a risk to health also needs its own report to FDA within 10 working days under 21 CFR 806.10, so the remedial action screen links to that record.
Complaint lifecycle
Six stages, each with an owner and a record. The aware date travels with the complaint from the first one.
AI triage limits
AI driven insights for medical device complaint handling are useful for sorting and grouping. They never decide whether a complaint is reportable.
Example: Sort a free-text complaint into categories and problem codes
Approach: AI proposes; the complaint handler accepts or edits, and both versions are stored
Example: Find similar complaints across lots, sites and channels
Approach: AI groups; a quality engineer confirms before a group is used to skip a repeat investigation
Example: Spot words that suggest injury or death in a long narrative
Approach: A flag can only raise priority, never lower it, and trained staff still read the complaint
Example: Draft the event narrative for the report
Approach: A regulatory specialist edits and approves every word before submission
Example: Decide reportable or not reportable, or close a complaint
Approach: People only, with their reasoning recorded in the MDR event file
SaMD complaints
Optimizing SaMD complaint handling means catching complaints that never arrive as a returned device.
Modules we scope one by one, so a first release can cover one product line and still meet every rule that applies to it.
Forms for call center, field and web complaints with the QMSR record fields enforced, plus connectors for support and app store channels.
Every complaint carries its aware date, and the 30-day and 5-day clocks show on each screen and in daily alerts.
Device history, lot and UDI lookup, returned device analysis, attachments and a link to earlier investigations of similar complaints.
Questions built from the decision table, a required reasoning field and the name and time of the person who decided.
Report content assembled from the complaint record, submitted electronically, with acknowledgments and supplemental reports tracked.
Category suggestions, injury flags and similar-complaint groups, each confirmed by a person and logged.
Deliberations, reports and acknowledgments kept together for each event and retained for the required period.
Each complaint links to corrective action and trend analysis in your quality management system.
Health Insurance Portability and Accountability Act
Protect PHI with privacy-first architecture, encrypted storage and transmission, strict access controls, and traceable audit logs.
General Data Protection Regulation
Implement lawful consent flows, data minimization, retention controls, and secure processing for sensitive health data.
Fast Healthcare Interoperability Resources
Enable standardized health data exchange across apps, care teams, and systems through standards-based FHIR APIs.
Health Level Seven International
Support enterprise-grade interoperability with HL7-based integrations for records, events, and clinical messaging workflows.
Health Information Trust Alliance
Align security programs to healthcare-specific control and risk management practices trusted by providers and ecosystem partners.
Health Information Technology for Economic and Clinical Health Act
Design with breach notification readiness, digital record safeguards, and operational controls that support regulated care programs.
FDA Software as a Medical Device
Plan software quality, traceability, and documentation pathways for products that may require SaMD review and submission.
Medical Device Regulation (European Union)
Prepare EU market-ready processes for risk classification, evidence tracking, and lifecycle governance under MDR expectations.
Substance Abuse and Mental Health Services Administration
Apply confidentiality controls and consent-aware sharing models for behavioral and mental health data experiences.
Standards and rules
The regulations and standards behind the decision table and the complaint record.
Rules the reportability screen and complaint record are built from, each linked to its primary source.
A device manufacturer must report to FDA no later than 30 calendar days after becoming aware of information, from any source, that reasonably suggests a device it markets may have caused or contributed to a death or serious injury, or has malfunctioned in a way that would likely cause one if it recurred.
Source: 21 CFR 803.50(a) (eCFR)A manufacturer must submit a 5-day report no later than 5 work days after becoming aware that a reportable event needs remedial action to prevent an unreasonable risk of substantial harm to public health, or after FDA asks for one in writing.
Source: 21 CFR 803.53 (eCFR)Manufacturers and importers must submit initial, supplemental and follow-up medical device reports to FDA in an electronic format that FDA can process, review and archive.
Source: 21 CFR 803.12(a) (eCFR)MDR event files must hold the documentation of the deliberations and decision-making used to decide whether an event was reportable, and a manufacturer keeps each file for 2 years from the event or the expected life of the device, whichever is greater.
Source: 21 CFR 803.18(b) and (c) (eCFR)Under the Quality Management System Regulation, complaint records for reportable or investigated complaints must include the device name, date received, any UDI or UPC, the complainant's contact details, the nature of the complaint, any correction or corrective action, and any reply to the complainant.
Source: 21 CFR 820.35(a) (eCFR)Who it is for
Teams that need complaint handling built around their products and channels.
Complaints, investigations and reports moved from shared files into one system with clocks.
Intake from app stores and support tools, with software version and telemetry attached to each complaint.
A complaint process ready before the first commercial launch, sized for a small quality team.
Complaint intake and report routing for importers, who carry their own reporting duties.
Category and grouping suggestions added to an existing complaint system, with people confirming each one.
Scope and cost
We do not publish prices. These are the things that change the size of a build, and we price them on the scoping call.
Call center, field service, web, email, support tools and app stores each add a connector and tests.
How many device families, and whether software products sit beside hardware.
Your QMS, ERP for lot and UDI data, CRM and service systems.
How many reports you file, which shapes how much of the eMDR preparation is automated.
Whether AI suggestions are in the first release, and the review and testing they need.
Computer software assurance and Part 11 evidence your quality team needs before go-live.
Three stages, each with a clear output.
We map your channels, products, current complaint flow and where reports have run close to their clocks.
ScopeIntake, investigation, the reportability decision and the event file for one product line, with validation evidence.
BuildMore channels, products and AI triage, plus support under your change control procedure.
Scale